
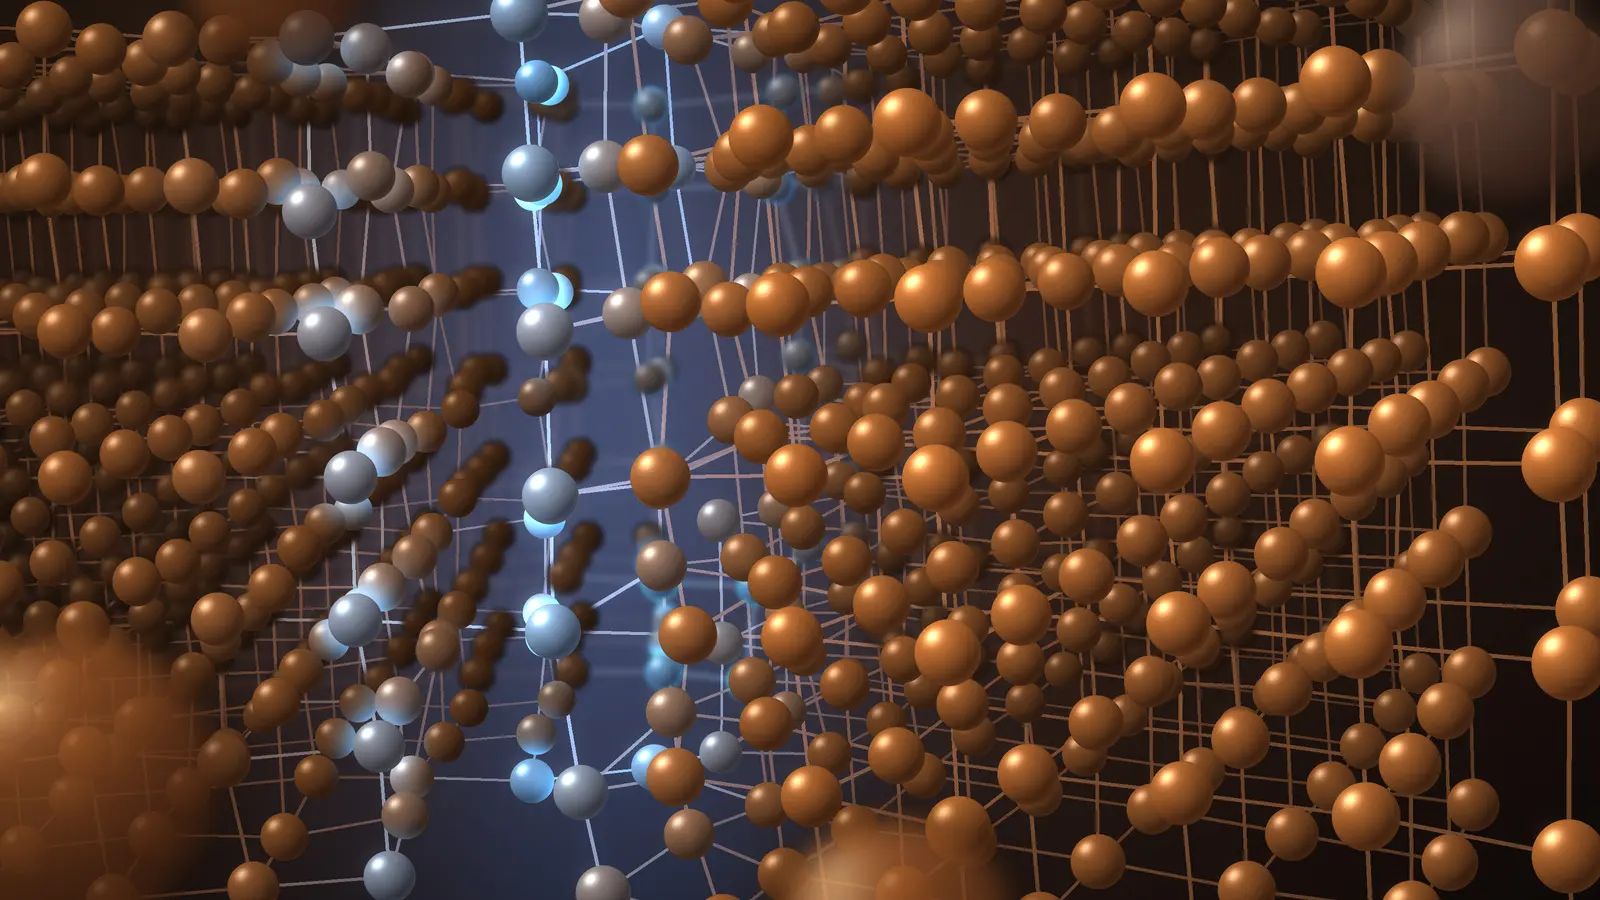
Electron-phonon interactions play a central role in many condensed matter phenomena, including electrical transport, superconductivity, polaron formation, and ultrafast carrier dynamics. However, research has traditionally focused on linear, first-order electron-phonon couplings. Second- and higher-order nonlinear interactions are often neglected because calculating them accurately in real materials can be computationally demanding, while existing methods have struggled to provide both diagonal and off-diagonal coupling matrix elements.
Researchers at The University of Texas at Austin and the Oden Institute have developed a theoretical framework and computational method for calculating nonlinear electron-phonon interactions of any order in real materials. The approach combines calculations of electron wavefunctions within a crystal unit cell with calculations of phonon perturbations in larger supercells, providing a systematically improvable workflow that can be used with different electronic-structure methods and Wannier interpolation.
High-Accuracy Calculation of Electron-Phonon Couplings
The new framework provides accurate calculations of both linear and nonlinear electron-phonon coupling strengths. For linear coupling matrix elements, the method achieved a mean absolute error below one milli-electronvolt, compared with errors exceeding 10 meV reported for earlier approaches that relied on approximations.
The researchers also demonstrated that the method can accurately calculate diagonal and off-diagonal nonlinear coupling matrix elements, which have previously been difficult or prohibitively expensive to obtain. By combining unit-cell and supercell calculations with Wannier interpolation, the workflow maintains high accuracy while allowing relatively large atomic displacement parameters in finite-difference calculations.
Tests on diamond, lithium fluoride (LiF), and graphite showed errors below 0.7% at a displacement of 0.01 Å. The three materials were selected to represent different classes of systems: diamond as a nonpolar semiconductor, LiF as a polar semiconductor, and graphite as a metal. This demonstrates the broad applicability of the computational approach.
The workflow begins with density functional theory (DFT) calculations of electronic states in the crystal unit cell and then incorporates phonon perturbations calculated using larger supercells. Vibrational modes and frequencies can be obtained using density functional perturbation theory (DFPT) or finite-difference methods.
Computing Second-Order Electron-Phonon Interactions
A key result of the work is a computationally tractable expression for second-order electron-phonon coupling matrix elements that can be evaluated using first-principles calculations. These matrix elements describe processes in which an electron changes from one electronic state to another while interacting with two phonons.
The framework calculates matrix elements of the form (g_{mn\nu\nu’}(\mathbf{k},\mathbf{q},\mathbf{q}’)), describing transitions between electronic states accompanied by the absorption or emission of phonons from branches (\nu) and (\nu’) with wavevectors (\mathbf{q}) and (\mathbf{q}’).
Accurately evaluating these quantities is challenging because the relevant expressions involve integrations over a Born-von Karman supercell, while the electronic wavefunctions are naturally described within the unit cell. The new method addresses this difficulty by combining the complementary advantages of unit-cell electronic calculations and supercell phonon perturbations.
The framework can also be systematically improved and used with different exchange-correlation functionals. Its compatibility with Wannier-Fourier interpolation further allows the calculated coupling coefficients to be efficiently transferred across momentum space.
Nonlinear Couplings Improve Polaron Calculations
The researchers found that second-order interactions can have a substantial impact on calculations of polaron properties. In particular, the Debye-Waller self-energy associated with quadratic electron-phonon terms can be comparable in magnitude to the Fan-Migdal self-energy arising from linear couplings.
This finding is important because both contributions influence calculations of temperature-dependent electronic structures within Allen-Heine theory. The researchers therefore extended ab initio polaron equations to incorporate second-order electron-phonon couplings.
The resulting calculations show that nonlinear interactions are important for obtaining quantitatively accurate polaron formation energies and hopping barriers. Including these contributions could improve theoretical descriptions of how charge carriers interact with atomic vibrations and move through materials.
Broader Applications
The new framework provides a route toward calculating electron-phonon interactions beyond the conventional linear approximation. Because the method can be generalized to interactions of higher order, it could support investigations of systems where nonlinear and anharmonic effects play an important role.
Potential applications include phonon-mediated superconductivity, charge transport, polaron physics, ultrafast carrier dynamics, and excited-state processes. More accurate descriptions of electron-phonon interactions could also help researchers understand energy and charge transfer in materials used for solar cells and other electronic devices.
By providing a practical first-principles method for calculating nonlinear electron-phonon couplings, the work extends computational tools for studying how electrons and atomic vibrations interact in real materials.
👉 More information
🗞 Nonlinear electron-phonon interactions from first principles
✍️ Zhenbang Dai and Feliciano Giustino
🧠 DOI: https://doi.org/10.1103/q5sl-wtlp
See today’s quantum computing news on Quantum Zeitgeist for the latest breakthroughs in qubits, hardware, algorithms, and industry deals.