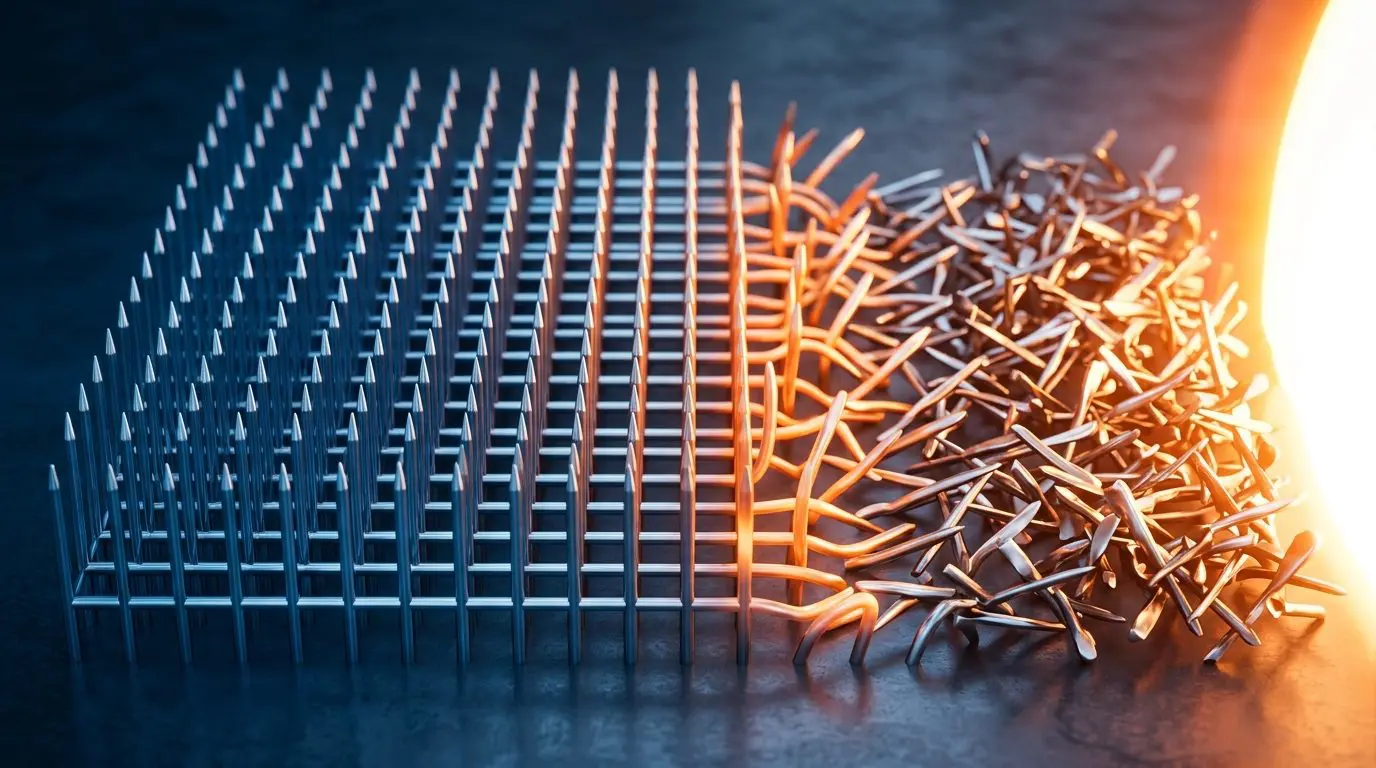
Researchers from the University of South Florida, Georgia Institute of Technology, Trinity College Dublin, and Queen’s University Belfast have developed a new approach to modeling the phase transitions in ferroelectric and antiferroelectric materials. The team demonstrates that these transitions, triggered by both temperature and electric fields and characterized by hysteresis loops, can be more accurately modeled using quantum mechanics rather than traditional classical methods.
This work overcomes limitations of existing simulations and highlights the fundamental role of quantum effects in materials long considered purely classical. The researchers anticipate this framework will extend to modeling a wide range of phase transitions, including magnetic and electronic phenomena.
Relaxation Modeling of Ferroelectric and Antiferroelectric Transitions
Simulations utilizing a frequency of 1.7 terahertz and time steps of 0.01 femtoseconds successfully reproduced hysteresis loops across a range of antiferroelectric materials, demonstrating the model’s capacity to represent complex material behaviors. This achievement departs from conventional modeling techniques that often rely on classical interpretations of phase transitions in ferroelectric and antiferroelectric substances, instead prioritizing a quantum mechanical treatment of relaxational processes.
The approach centers on describing a quantum state evolving in time, governed by a Hamiltonian incorporating momentum and potential energy, represented mathematically as |ψ(t)⟩ =, where |⟩ and are the eigenstates and eigenvalues of the Hamiltonian. The framework’s success in semi-quantitatively reproducing hysteresis loops provides compelling evidence that quantum mechanics and relaxation dynamics are not merely supplemental to, but essential components of, these phase transitions.
Researchers employed a Ground-State Relaxation model, tracking the time evolution of a quantum state |ψ(t)⟩ influenced by a time-independent Hamiltonian, to simulate the behavior of these materials under varying conditions. Further simulations, conducted with a frequency of 0.17 terahertz and 0.1 femtosecond time steps, expanded the model’s validation to additional antiferroelectric materials, reinforcing its broad applicability. This unconventional methodology offers a powerful alternative to traditional modeling, which often assumes Arrhenius-type activation processes occurring within a classical regime.
The work demonstrates that by framing phase transitions as relaxational processes requiring quantum mechanical consideration, accurate and efficient first-principles simulations become attainable. “Based on our results, we conclude that treating (anti)ferroelectric phase transitions as relaxational processes within a quantum mechanical framework offers an unconventional yet powerful approach to their modeling and understanding,” the researchers state, highlighting the significance of this shift in perspective.
Quantum Mechanical Approach Overcomes Classical Limitations
Conventional simulations of ferroelectric and antiferroelectric materials often treat phase transitions as activated processes occurring within a classical framework, yet recent work demonstrates a viable alternative rooted in quantum mechanics and relaxational dynamics. This shift in perspective allows for more efficient, first-principles simulations of these transitions, addressing limitations inherent in earlier modeling techniques.
The research extends beyond simply replicating observed phenomena; it reveals the significant role quantum effects play in transitions previously considered purely classical. Path integral techniques, previously applied to strontium titanate and barium titanate, have demonstrated that quantum phenomena can substantially alter transition temperatures, by as much as 50 Kelvin in barium titanate, and enhance dielectric and piezoelectric responses.
These findings suggest that even at room temperature, quantum effects are not negligible in these materials, challenging long-held assumptions about their behavior. Further simulations, employing a frequency of 0.17 terahertz and time steps of 0.1 femtoseconds, also confirmed these results. The implications of this new approach extend far beyond ferroelectrics and antiferroelectrics, potentially impacting the modeling of a diverse range of phase transitions.
Researchers anticipate applicability to magnetic, elastic, and multiferroic transitions, as well as phenomena like quantum tunneling and chemical reaction rates. This framework not only provides a deeper fundamental understanding but also promises more reliable property predictions for these important materials, opening avenues for advanced materials design and application.
Double-Well and Triple-Well Free Energy Profiles Define Phase States
The shapes of free energy profiles, specifically, double-well forms for ferroelectrics and triple-well forms for antiferroelectrics, dictate how these materials change phase under varying conditions, as revealed by new computational modeling. These profiles, representing energy as a function of polarization, are fundamental to understanding the behavior of materials used in memory devices, sensors, and high-energy capacitors. Traditional interpretations of these transitions relied on the assumption that they were activated processes, with transition rates proportional to the exponential of negative activation energy divided by Boltzmann’s constant times temperature.
This approach directly links to the equation of state, derived by setting the derivative of free energy, modified to include electric field and volume, to zero. Avoiding thermodynamically unstable branches then generates the characteristic hysteresis loops observed in these materials.
This method allows for quantifying the electric fields needed to overcome energy barriers, which is an important step in predicting material response. The model’s applicability extends beyond simply replicating observed behavior; it provides a means to probe quantum-mechanical features underlying a broad range of phase transitions.
Calculations were performed using the Vienna Ab initio Simulation Package (VASP), focusing on energy profiles to establish the foundation for this new approach. The framework also directly addresses phase transitions driven by electronic degrees of freedom, such as proton-ordering and hopping ferroelectrics, broadening its potential impact. the modeling incorporates a summation over states, replacing it with a density of states for the heat bath, a technique that refines the calculation of transition rates.
DFT Simulations Reveal Polarization Behavior in PbTiO3 and PbZrO3
Calculations of energy per formula unit reveal distinct behaviors in lead titanate (PbTiO3) and lead zirconate (PbZrO3), materials central to modern electronics and sensing technologies. Density functional theory simulations demonstrate that the energy landscape of antiferroelectric PbZrO3 exhibits features not fully captured by classical modeling approaches, suggesting a more nuanced relationship between polarization and applied electric fields than previously understood. These simulations, alongside those for PbTiO3, provide a detailed view of the energy changes accompanying phase transitions, offering insights into the stability of metastable states.
The simulations detail how the transition rate between these states is linked to the height of the energy barrier, a concept rooted in activated processes where lower barriers facilitate faster transitions. This approach allows for the computation of polarization dynamics under applied electric fields, as shown in simulations of PbZrO3 transitioning from a metastable to a ground state under a 400 kV/cm field.
The resulting hysteresis loops, a hallmark of ferroelectric and antiferroelectric materials, emerge naturally from this framework. Beyond replicating observed behavior, the modeling provides quantifiable data on the energy barriers influencing phase changes.
The work details specific material properties, such as the soft mode frequency of 1 THz, alongside parameters which contribute to a more complete understanding of the underlying physics governing these transitions and offer a pathway to predict material responses under varying conditions. The framework’s applicability extends to materials like AlN and HfO2, as evidenced by computed energy landscapes for these compounds, suggesting a broad potential for materials design and optimization.
Arrhenius-Type Transition Rate Fails at High Energy Barriers
Density functional theory simulations reveal that conventional modeling of phase transitions in materials like barium titanate relies on an inaccurate assumption regarding energy barriers. Specifically, the widely used Arrhenius-type transition rate, which predicts how quickly a material shifts between states, fails when the energy barrier separating those states is particularly high. This failure manifests as unrealistically low transition rates, and consequently, extremely long lifetimes for metastable states, a discrepancy observed even with refinements to the model incorporating frequency and temperature dependencies.
Calculations show that for a typical soft mode frequency of 1 terahertz, the transition rate at room temperature and an experimental electric field of 315 kilovolts per centimeter is 2.0 × 10-40, a value deemed unphysical. However, this framework struggles to accurately represent the dynamics when the energy barrier is substantial, leading to an overestimation of the required electric field to induce a phase change.
Previous attempts to correct for this have focused on improving the accuracy of the density functional theory calculations themselves, aiming to better capture the transition states and associated barriers. The current work proposes a different solution, shifting the underlying assumptions about how these transitions occur. Instead of viewing the phase changes as simply overcoming an energy barrier, the research team proposes a relaxational process requiring a quantum mechanical treatment.
Equation of State Modeling and Electric Field Influence on Polarization
Simulations of lead zirconate (PbZrO3) reveal a computed energy landscape dependent on temperature, providing insight into antiferroelectric behavior. Black and green dots within the simulations delineate metastable and ground states, respectively, mirroring experimental polarization data obtained through applied electric fields. These computational results, alongside similar analyses of barium titanate and other materials, utilize parameters like a vibrational frequency, denoted as ν, typically around 0.17 terahertz, though varying up to 1.7 terahertz for titanium dioxide.
The time step, Δt, remained at 0.01 femtoseconds for simulations involving titanium dioxide, cesium germanium bromide, aluminum nitride, and hafnium dioxide. Further refinement of these simulations incorporates a scaling factor, , which ranges from 0.007 for aluminum nitride to 0.045 for lead zirconate, influencing the modeled polarization response to electric fields across a temperature spectrum from 10 to 1000 Kelvin.
The parameters used in these calculations are not merely inputs, but define the precision with which the model can replicate observed material behavior under diverse conditions, offering a detailed view of how these materials respond to external stimuli. These detailed simulations, focusing on parameters like vibrational frequency, time step, and scaling factor, provide a granular understanding of how different materials exhibit polarization changes with temperature and electric field. The consistency of the time step across some materials, coupled with the variation in scaling factors, suggests a nuanced approach to modeling complex phenomena.
Zero-Kelvin Behavior Predicts Mixed Loops in PbZrO3
Simulations reveal lead zirconate exhibits unexpected antiferroelectric-like behavior at absolute zero, displaying double hysteresis loops alongside spontaneous polarization, a stark contrast to experimental measurements of the material. The discrepancy arises because traditional modeling approaches often fail to capture quantum effects crucial to understanding phase transitions in these materials, and the simulations used an applied electric field of 400 kV/cm. Researchers constructed a distortion path between crystal structures, revealing a predicted response not fully mirrored results.
Computed polarization values are derived from the expectation value of the polarization operator, offering a theoretical framework for predicting material responses to external stimuli. The simulations, performed with a time step of 1 femtosecond and an AC field frequency of 1.7 THz, successfully reproduce the trend of larger coercive fields in materials like aluminum nitride and hafnium dioxide compared to oxide ferroelectrics such as lead titanate, validating its predictive power across a range of compositions.
Notably, the model predicts a larger coercive field for cesium germanium bromide than for lead titanate, potentially linked to the specific exchange-correlation functional employed in the calculations. The team utilized a distortion path, and could not use LDA because it incorrectly predicted the stable phase, disagreeing with experimental data. The paper reports, highlighting the framework’s ability to align with established material properties. While the model predicts coercive fields of approximately 3018 kV/cm for ferroelectrics, experimental measurements report values around 175 kV/cm, indicating a significant divergence between theory and observation that warrants further investigation.
Unphysical Coercive Fields Highlight Need Beyond Transition State Theory
Calculated energy barriers in traditional models often fail to align with observed transition rates; simulations using a 1 terahertz soft mode frequency predict a transition rate of 2.0 × 10⁻⁴⁰ s⁻¹ at room temperature and an applied field of 315 kV/cm, a value demonstrably unphysical. This discrepancy stems from the reliance on Arrhenius-type processes, which assume energy barriers significantly exceed thermal energy, and a classical treatment of the transitions themselves.
The research demonstrates that a relaxational model, incorporating quantum mechanical treatment, offers a more accurate framework for understanding these phenomena. Previous attempts to refine the traditional model, such as incorporating frequency- and temperature-dependent transition rates, still yielded coercive fields higher than those measured experimentally, suggesting fundamental limitations within the existing approach.
The team addressed this by formulating a model where population changes between energy levels are governed by both coherent evolution and relaxation. The model’s success with ferroelectrics, correctly predicting coercive fields that exceed experimental ones (~175 kV/cm), further validates its broader applicability beyond simple ferroelectric materials.
See today’s quantum computing news on Quantum Zeitgeist for the latest breakthroughs in qubits, hardware, algorithms, and industry deals.